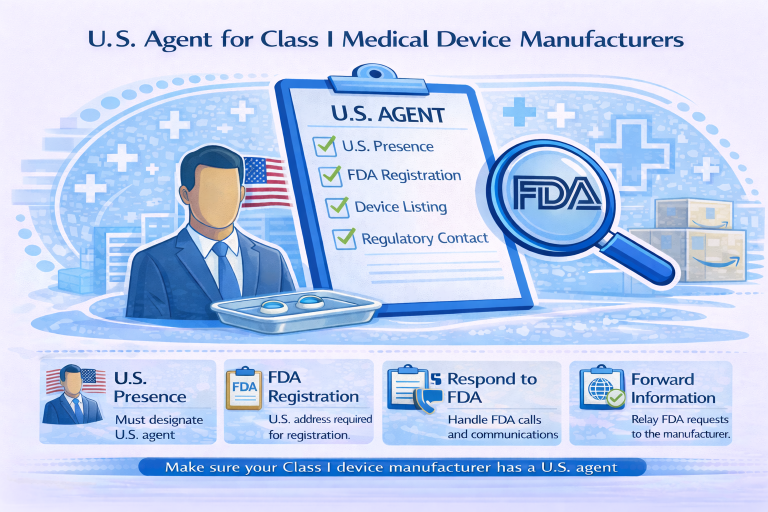
U.S. Agent for Class I Medical Device Manufacturers
"510(k)-exempt" is the single most misunderstood phrase in Class I compliance. Here's what it actually means — and everything it doesn't.
Key Takeaways
- Foreign Class I makers need a U.S. Agent (21 CFR 807.40) — plus an Official Correspondent.
- 510(k)-exempt ≠ requirement-free: registration, listing, QMSR, labeling, UDI, and MDR all still apply.
- Not every Class I device is exempt — exemption limitations can still trigger a 510(k).
- Annual fee: $11,423 (FY2026), paid Oct 1–Dec 31; reset each fiscal year; no separate listing fee.
- Miss Dec 31 → "not duly registered" and devices can be treated as misbranded.
- QMSR is now in effect (Feb 2, 2026) — a quality system consistent with ISO 13485 is expected where applicable.
What "Class I" Means
Class I devices are the lowest-risk category — think bandages, exam gloves, manual surgical instruments, and many hospital supplies. Because the risk is low, most are subject only to "general controls" and most are exempt from 510(k) premarket notification. But "low risk" doesn't mean "unregulated": general controls still include registration, listing, labeling, and quality obligations.
What a Foreign Class I Manufacturer Must Do
Here's the complete obligation set — the U.S. Agent is just the entry point:
| Requirement | What it involves | Applies? |
|---|---|---|
| Establishment registration | Register in FURLS/DRLM annually; pay the fee (21 CFR 807) | Required |
| Device listing | List each device with product code and activities | Required |
| U.S. Agent | Domestic FDA contact for the foreign establishment (807.40) | Required |
| Official Correspondent | Handles registration/listing communications | Required |
| QMSR (quality system) | 21 CFR 820 / ISO 13485:2016 — effective Feb 2, 2026 | Where applicable |
| Labeling & UDI | Device labeling (21 CFR 801) and Unique Device Identifier | Required |
| Medical Device Reporting | Report certain adverse events (21 CFR 803) | Required |
| 510(k) premarket notification | Only if the device isn't exempt / exceeds exemption limits | Often exempt |
The Annual Fee — and the Deadline That Bites
The fee is flat — it applies to a small Class I manufacturer the same as a large one. Registration and payment must be completed in the October 1–December 31 window each year. Miss it, and your establishment is "not duly registered" for the new fiscal year — which can make your devices misbranded, exposing them to import detention and warning letters.
U.S. Agent vs. Official Correspondent — and the Limits
U.S. Agent
FDA's U.S. contact for the foreign establishment: assists communications, answers questions about your devices, and helps schedule inspections. If FDA can't reach you directly, delivering to the agent counts as delivering to you.
Official Correspondent
Facilitates registration and listing communications and can submit that data in FURLS. A foreign establishment has both roles — the same person can fill them, but they're distinct.
How to Get Your Class I Device Compliant
Confirm classification & exemption limits
Verify your product code and Class I status, and check the limitation section to confirm the device is truly 510(k)-exempt.
Designate a U.S. Agent & Official Correspondent
Both are needed for a foreign establishment; the same provider can serve in both roles.
Register & pay the fee
Register in FURLS/DRLM and pay the annual fee within the Oct 1–Dec 31 window.
List your devices
List each Class I device with its product code and activities — required for legal U.S. distribution.
Meet quality, labeling & reporting rules
Comply with QMSR, labeling and UDI, and medical device reporting where applicable.
Renew & maintain annually
Re-register and pay each fiscal year, keep listings current, and update your U.S. Agent as needed to stay duly registered.
U.S. Agent + Full Class I Compliance
FDA Registration Assistance serves as your U.S. Agent and Official Correspondent, completes establishment registration and device listing, tracks the fee and renewal window, and helps align QMSR, labeling, UDI, and reporting — so a 510(k)-exempt device doesn't get tripped up by the requirements that still apply.
Contact us at info@fdaregistrationassistance.com or call +1 (928) 275-8333. Trusted by device makers in 135+ countries, with 15+ years of FDA experience.
Frequently Asked Questions — Class I Devices & U.S. Agent
1. Do Class I device manufacturers need a U.S. Agent?
Foreign ones do. Any non-U.S. establishment making Class I devices for the U.S. market must designate a U.S. Agent as part of registration (21 CFR 807.40). Domestic U.S. makers don't. It's one of several requirements, not the only one.
2. Class I devices are 510(k)-exempt — doesn't that mean fewer rules?
Being 510(k)-exempt only removes premarket notification. It doesn't exempt registration, listing, the U.S. Agent requirement, QMSR, labeling, UDI, or MDR. "Exempt from 510(k)" is not "exempt from FDA."
3. Is every Class I device actually 510(k)-exempt?
Most are, not all. Even a listed exempt product code has limitations (the ".9" sections) tied to technology, intended use, or features. Exceed them and a 510(k) may be required — so confirm the code and its limits first.
4. What's the annual establishment registration fee?
For FY2026 (Oct 1, 2025–Sep 30, 2026) it's $11,423, up ~23% year over year. FDA resets it each fiscal year. It's flat — applying to Class I too — and there's no separate FDA listing fee.
5. Is there fee relief for small manufacturers?
For FY2026, FDA introduced a limited waiver (excluding initial registration) for certain small businesses that qualify through the Small Business Determination program and show financial hardship. It's not automatic. Historically there was no waiver for this fee.
6. When is the fee due?
Registration and payment must be completed in the Oct 1–Dec 31 window each year. Distributing after Dec 31 without completing it leaves you not duly registered, and devices can be treated as misbranded.
7. What if I miss the December 31 deadline?
FDA treats the establishment as not duly registered for the new fiscal year. Devices from an unregistered establishment can be deemed misbranded, risking import detention and warning letters — FDA has issued such letters for missed registration and listing.
8. What's an Official Correspondent, and do I need one?
For devices, the Official Correspondent facilitates registration and listing communications and can submit that data in FURLS. A foreign establishment has both a U.S. Agent and an Official Correspondent; one person can fill both, but they're distinct roles.
9. What does the U.S. Agent do for a device establishment?
It's FDA's U.S. contact for the foreign establishment — assisting communications, answering questions about your devices, and helping schedule inspections. If FDA can't reach you directly, delivering to the agent counts as delivering to you.
10. Is the U.S. Agent responsible for my MDR or 510(k)?
No. The U.S. Agent has no responsibility for medical device reporting (21 CFR 803) or submitting 510(k)s. Those stay with the manufacturer — the agent is a liaison, not a stand-in for your obligations.
11. What is QMSR, and does it apply to Class I?
QMSR is the updated 21 CFR 820, incorporating ISO 13485:2016, effective Feb 2, 2026, replacing the old Quality System Regulation. It applies to device makers where applicable — including many Class I — so a quality system consistent with the standard is expected even for exempt devices.
12. Do Class I devices need a UDI?
Generally yes, unless specifically excepted. UDI labeling applies across classes and Class I compliance dates have passed, so most Class I devices need a UDI on labels and packaging. Confirm your specific device for any exception.
13. Does registration mean my device is FDA approved or cleared?
No. Registration and listing only identify your facility and products to FDA — not approval, clearance, or endorsement. FDA issues no registration "certificate," and calling a registered Class I device "FDA approved" is inaccurate.
14. Can my importer or distributor be my U.S. Agent?
Possibly, but often not ideal — a distributor has its own interests, and if the relationship ends you can lose your agent. The initial importer is also a separate registrant with its own duties; being your importer doesn't make it your U.S. Agent by default.
15. Does each facility need its own registration and U.S. Agent?
Yes. Each establishment registers independently by role, and each foreign establishment designates one U.S. Agent. A contract manufacturer's registration doesn't cover yours — you register separately.
16. Can I change my U.S. Agent later?
Yes — update the agent in your registration, and the new agent confirms consent with FDA. Changes to the agent's name, address, or phone must be reported within 10 business days. See our switching guide.
17. How does FDA Registration Assistance help?
It serves as your U.S. Agent and Official Correspondent, completes registration and listing, tracks the fee and renewal window, and helps align QMSR, labeling, UDI, and reporting — so an exempt device isn't tripped up by what still applies.
18. How do I get started?
Contact FDA Registration Assistance at info@fdaregistrationassistance.com or call +1 (928) 275-8333 with your device type and product code for a clear compliance plan.