FDA & Dietary Supplements: What's Actually Required
"No approval needed" is the part everyone hears — and the part that trips companies up. The obligations are real; they just apply differently than for drugs.
Key Takeaways
- Regulated under DSHEA as food — no premarket approval, but real rules.
- Facility registers with FDA; foreign facilities add a U.S. Agent.
- cGMP is mandatory — 21 CFR Part 111, enforced since 2010.
- Labels need a Supplement Facts panel (21 CFR 101.36) and honest claims.
- Structure/function claims yes, disease claims no — disease claims can make it a drug.
- New Dietary Ingredients need a 75-day notification (21 CFR 190.6).
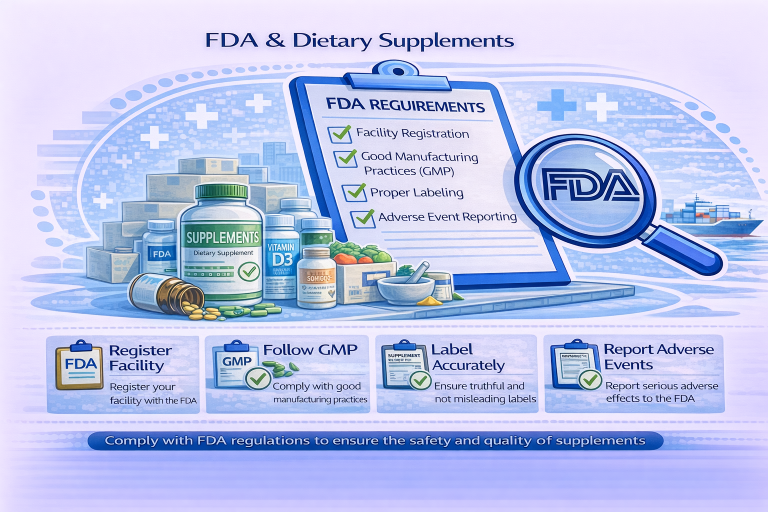
Six Things FDA Expects
| Requirement | What it means | Where it lives |
|---|---|---|
| Facility registration | The maker/packer/holder registers as a food facility; foreign facilities add a U.S. Agent | 21 CFR Part 1, Subpart H |
| cGMP | Identity/purity testing, process controls, batch records | 21 CFR Part 111 |
| Labeling | Supplement Facts panel, identity, net quantity, ingredients, firm name/address | 21 CFR 101.36 |
| Claims | Structure/function allowed with disclaimer; disease claims prohibited | FD&C Act / DSHEA |
| New Dietary Ingredients | 75-day premarket notification with safety data | 21 CFR 190.6 |
| Adverse events | Receive and report serious adverse events; label contact | AER Act (2006) |
Structure/Function vs. Disease Claims
The single most expensive error in this category is claiming too much. The line is simple to state and easy to cross:
Allowed: structure/function
- "Supports normal immune function"
- "Helps maintain healthy bones"
- Must carry the DSHEA disclaimer
- Notify FDA within 30 days of first use
Not allowed: disease claims
- "Treats arthritis" / "cures colds"
- "Lowers blood pressure"
- "Prevents cancer"
- Can reclassify the product as an unapproved drug
Two Places Companies Get Caught
New Dietary Ingredients (NDI)
If an ingredient wasn't marketed in the U.S. before October 15, 1994, it may be a New Dietary Ingredient — requiring a notification with safety data at least 75 days before marketing (21 CFR 190.6), unless it's been in the food supply unaltered. FDA's acknowledgment isn't a safety finding or approval.
Importing supplements
Imported supplements need a registered facility, a U.S. Agent, and prior notice per shipment — and the U.S. importer carries FSVP duties (modified for supplements). Gaps here mean held or refused entries.
Path to a Compliant Launch
Register the facility
The maker/packer/holder registers with FDA; foreign facilities add a U.S. Agent.
Manufacture under cGMP
Follow 21 CFR Part 111 — identity testing, process controls, batch records.
Check ingredients for NDI status
Flag any ingredient new since Oct 15, 1994, and notify FDA 75 days ahead if needed.
Build a compliant label
Supplement Facts panel, identity, net quantity, ingredients, firm info — and lawful claims.
Set up adverse event reporting
Put a system in place to receive and report serious adverse events.
Handle imports
Registration, U.S. Agent, prior notice, and FSVP all in place before the first shipment.
Get Your Supplement to Market the Right Way
FDA Registration Assistance registers supplement facilities, serves as U.S. Agent, reviews Supplement Facts labels and claims, and helps assess NDI status and import requirements — so the pieces line up before you sell.
Contact us at info@fdaregistrationassistance.com or call +1 (928) 275-8333. Trusted by companies across 135+ countries, with 15+ years of experience.
Frequently Asked Questions
1. Are dietary supplements regulated by the FDA?
Yes — under DSHEA (1994), which treats them as a category of food, not drugs. They don't need approval before sale, but must meet requirements for manufacturing, labeling, claims, and safety, with FDA oversight mainly after market.
2. Do supplements need FDA approval before being sold?
No — unlike drugs, there's no premarket approval. The manufacturer is responsible for a safe, properly labeled product; FDA can act against unsafe or misbranded supplements once they're on the market.
3. Do I need FDA facility registration for supplements?
Yes — the facility that makes, packs, or holds the supplement registers as a food facility, and foreign facilities add a U.S. Agent. Registration identifies the facility; it isn't product approval.
4. What is cGMP for supplements?
The dietary supplement cGMP rule, 21 CFR Part 111, requires identity and purity testing, process controls, qualified staff, and batch records so products match their labels. It's been enforced since 2010.
5. What has to be on a supplement label?
A statement of identity, net quantity, a Supplement Facts panel (21 CFR 101.36), a full ingredient list, the firm's name and address, and any required disclaimer. Labeling errors are a leading reason supplements draw FDA attention.
6. What claims can a supplement make?
Structure/function claims (e.g., "supports normal immune function"), but not disease claims. Structure/function claims must carry the DSHEA disclaimer and be notified to FDA within 30 days of first marketing.
7. What happens if I make a disease claim?
Your product can be treated as an unapproved new drug — a serious problem and a common warning-letter trigger. See our pages on label warning letters and the supplement label crackdown.
8. What is a New Dietary Ingredient (NDI)?
An ingredient not marketed in the U.S. before October 15, 1994. If your product contains one, you generally submit an NDI notification with safety data at least 75 days before marketing (21 CFR 190.6), unless it's been in the food supply unaltered.
9. Does FDA "approve" an NDI notification?
No — FDA acknowledges and reviews it, but acknowledgment isn't a finding that the ingredient or product is safe or approved. You remain responsible for safety and lawful marketing.
10. Do I have to report adverse events?
Yes — companies must receive and report serious adverse events to FDA and include a contact for reports on the label. It's a standing obligation, not optional.
11. What about importing supplements?
Imports need a registered facility with a U.S. Agent, prior notice per shipment, and compliant labeling, plus the importer's FSVP obligations (modified for supplements). Gaps mean held or refused entries.
12. Is FDA changing how it regulates supplements?
FDA's supplement program has been modernizing its cGMP and labeling framework and working toward updated NDI guidance. A "mandatory product listing" has been proposed in Congress but isn't currently law — today's requirements apply.
13. Can I handle supplement compliance myself?
You can. Many use a firm because registration, cGMP, label review, NDI status, and imports are where costly mistakes happen. The requirements are the same either way.
14. How does FDA Registration Assistance help?
It registers supplement facilities, serves as U.S. Agent, reviews labels and claims, and helps assess NDI and import requirements for companies across 135+ countries. Contact info@fdaregistrationassistance.com or +1 (928) 275-8333.