FDA Compliance for Blood Pressure Monitors
Why BP monitors are Class II devices, the accuracy and safety standards, and why cuffless/wearable BP is still a device.
Key Takeaways
- BP monitor = Class II device — 21 CFR 870.1130, product code DXN.
- 510(k) required — not exempt; substantial equivalence to a predicate.
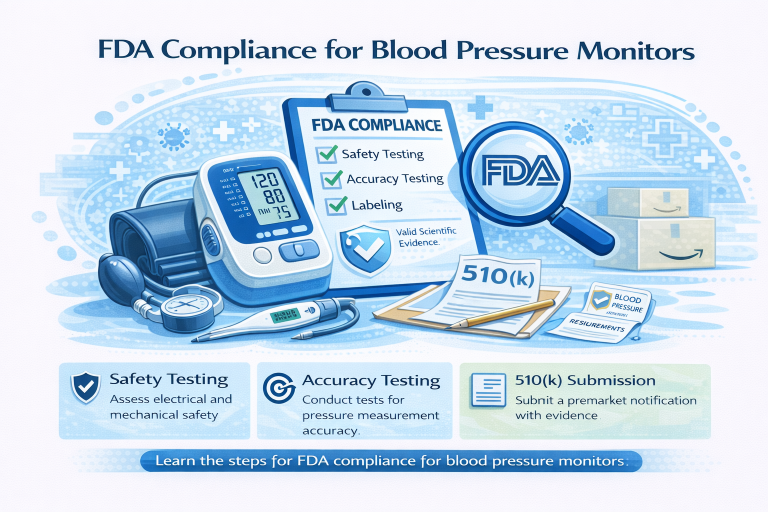
- Validate to ISO 81060-2 (accuracy), IEC 60601/EN 80601-2-30 (safety), ISO 10993 (cuff).
- Cuffless/wearable BP = still a device — FDA's Jan 2026 draft guidance uses ISO 81060-2.
- Disclaimers don't rescue a BP-measurement claim — intended use is judged from the whole presentation.
- "Cleared" ≠ "approved" — BP monitors are 510(k)-cleared.
Blood Pressure Monitors Are Class II Devices
A blood pressure monitor is a noninvasive blood pressure measurement system — a Class II device under 21 CFR 870.1130 (product code DXN, Circulatory System Devices Panel) — and it requires a 510(k) before it can be marketed:
- Covers automated/oscillometric monitors — measured at the arm, wrist, or finger; home consumer and clinical
- 510(k) required — BP monitors are not 510(k)-exempt; you demonstrate substantial equivalence to a predicate
- Ambulatory (ABPM) & multi-parameter monitors — the BP-measurement function is regulated the same way
- Cleared, not approved — a BP monitor is FDA-cleared via 510(k), not PMA-approved
Accuracy, Safety & Biocompatibility
| Standard | Covers |
|---|---|
| ISO 81060-2 | Clinical validation of measurement accuracy (automated/intermittent BP) — the key accuracy standard (successor to AAMI/ANSI SP10) |
| IEC 60601 / EN 80601-2-30 | Electrical safety and basic performance of the electronic monitor |
| ISO 10993 | Biocompatibility of the cuff and skin-contact materials |
Cuffless & Wearable BP Is Still a Device
Cuffless noninvasive BP measuring systems are generally Class II devices under the same regulation (21 CFR 870.1130, DXN), and FDA issued a January 2026 draft guidance specifically for cuffless BP devices — harmonizing testing with ISO 81060-2. So a smartwatch, band, or app that measures or reports blood pressure is performing a device function.
The Device Pathway for a BP Monitor
- Classification & 510(k) — confirm DXN/870.1130 and prepare the 510(k) with ISO 81060-2 accuracy validation and IEC 60601/EN 80601-2-30 safety testing
- Establishment registration & device listing — 21 CFR Part 807, via FURLS/DRLM
- Quality system — 21 CFR Part 820, now the QMSR
- Labeling & UDI — 21 CFR Part 801 and, where applicable, UDI (21 CFR Part 830); accuracy claims must match the validation
- Medical Device Reporting — 21 CFR Part 803
- U.S. Agent — for foreign establishments (21 CFR 807.40)
Common BP Monitor Mistakes
- Marketing a monitor without the required 510(k)
- Offering a cuffless/wearable BP feature as "wellness" without clearance
- Relying on a disclaimer instead of authorizing the device
- Inadequate ISO 81060-2 accuracy validation or overstated accuracy claims
- Calling it "FDA approved" (it's cleared); missing UDI
- Missing the U.S. Agent (foreign manufacturers)
Get Your Blood Pressure Monitor Compliant
FDA Registration Assistance confirms the classification (Class II, 21 CFR 870.1130, product code DXN), maps the 510(k) and standards strategy (ISO 81060-2 accuracy, IEC 60601/EN 80601-2-30 safety, ISO 10993 biocompatibility), and handles establishment registration and device listing, labeling/UDI, and U.S. Agent service for foreign makers — including for cuffless/wearable BP products where the device line is easy to cross.
Contact us at info@fdaregistrationassistance.com or call +1 (928) 275-8333 for a quote. An independent compliance firm serving 1,000+ companies across 135+ countries, with 15+ years of experience.
Frequently Asked Questions — Blood Pressure Monitors
1. Is a blood pressure monitor an FDA medical device?
Yes. A blood pressure monitor is a noninvasive blood pressure measurement system - a Class II medical device under 21 CFR 870.1130 (product code DXN, Circulatory System Devices Panel). It requires a 510(k) premarket notification (demonstrating substantial equivalence to a legally marketed predicate) before marketing. This covers automated/oscillometric monitors measured at the arm, wrist, or finger - both home consumer monitors and clinical monitors.
2. Do blood pressure monitors need a 510(k)?
Yes. As a Class II device, a blood pressure monitor generally requires a 510(k) clearance before it can be marketed - it isn't 510(k)-exempt. The 510(k) shows the device is substantially equivalent to a predicate and typically includes validation to the recognized performance and safety standards. Registration and listing don't substitute for the 510(k); you need the clearance for a BP monitor.
3. What standards does a blood pressure monitor need to meet?
The core ones are: ISO 81060-2, the clinical-validation standard for measurement accuracy of intermittent automated BP monitors (the key accuracy standard; the older AAMI/ANSI SP10 preceded it); IEC 60601 / EN 80601-2-30 for the electrical safety and basic performance of the electronic monitor; and ISO 10993 for biocompatibility of the cuff and skin-contact parts. A 510(k) for a BP monitor typically demonstrates conformance to these. The exact set depends on the device (cuff, wrist, cuffless, etc.).
4. What is ISO 81060-2 and why does it matter?
ISO 81060-2 is the international standard for clinically validating the accuracy of automated (intermittent) noninvasive blood pressure measurement - it defines the clinical study protocol and acceptance criteria a BP monitor must meet to demonstrate its readings are accurate. It's central to a BP monitor 510(k), because accuracy is the whole point of the device: inaccurate readings can lead to missed or mistaken hypertension management. FDA's recent cuffless-BP guidance also draws its methodology from ISO 81060-2.
5. Are cuffless or wearable blood pressure devices regulated?
Yes. Cuffless noninvasive blood pressure measuring systems are generally Class II devices under the same regulation (21 CFR 870.1130, product code DXN), and FDA issued a January 2026 draft guidance specifically for cuffless BP devices - harmonizing testing with ISO 81060-2 methodology. So a smartwatch, band, or app that measures or reports blood pressure is doing a device function and generally needs FDA authorization; it isn't automatically a "wellness" product just because it's a wearable.
6. Can I market a wearable BP feature as "wellness" to avoid FDA?
Be very careful - measuring blood pressure is a physiologic/diagnostic function, and FDA treats BP-measurement outputs as device functions. In 2025 FDA took action against a wearable company over a blood-pressure feature offered without clearance, inferring device/diagnostic intent from the company's own marketing (systolic/diastolic readings, "medical-grade" language, risk banding, and how users used it) under 21 CFR 801.4 - and disclaimers did not rescue the feature. So labeling a BP-measurement feature "wellness" or adding a disclaimer generally won't keep it out of device regulation.
7. Why don't disclaimers protect a BP-measurement feature?
Because FDA determines intended use from the objective evidence - your marketing, the way results are presented, and how consumers actually use the product - not just a disclaimer. If a product presents systolic/diastolic values, uses risk banding, or is marketed with "medical-grade" or health-management language, FDA can infer a device intent even without an explicit disease claim (21 CFR 801.4). A "not a medical device" disclaimer that contradicts the actual presentation doesn't cure the problem.
8. What registration does a BP monitor company need?
The device pathway: FDA establishment registration and device listing (21 CFR Part 807), the 510(k) clearance for the monitor (product code DXN), a compliant quality system (QMSR), device labeling and UDI where applicable, Medical Device Reporting, and a U.S. Agent if the establishment is foreign. Classification and the 510(k) (with the ISO 81060-2/IEC 60601 testing) are the front-end work; then registration/listing and ongoing compliance.
9. Is a blood pressure monitor "FDA approved"?
No - it's "FDA cleared," not "approved." BP monitors are Class II devices cleared through the 510(k) process (substantial equivalence), not premarket approval (PMA). Only high-risk Class III PMA devices are "FDA approved." Calling a BP monitor "FDA approved" is inaccurate; the correct description is FDA-cleared (via 510(k)), and your establishment is registered and the device listed.
10. Do wrist and finger monitors follow the same rules?
Yes - wrist and finger automated oscillometric monitors are also noninvasive blood pressure measurement systems (Class II, 21 CFR 870.1130, DXN) and require a 510(k) with accuracy validation. The measurement site affects the clinical validation and labeling, but the classification and pathway are the same. The device still has to demonstrate accuracy under the applicable standard for its site and method.
11. Do ambulatory blood pressure monitors (ABPM) need clearance too?
Yes. Ambulatory blood pressure monitors, which take repeated automated readings over 24 hours, are noninvasive BP measurement systems and are regulated as Class II devices requiring a 510(k) with appropriate validation. As with other BP monitors, the intended use (ambulatory, resting, sleep, etc.) must be clearly described and supported - FDA's guidance emphasizes clearly stating the measurement conditions the device is validated for.
12. Do foreign BP monitor manufacturers need to register?
Yes. A foreign manufacturer of blood pressure monitors must obtain the 510(k) clearance, complete FDA establishment registration and device listing, comply with the quality system and labeling/UDI requirements, and appoint a U.S. Agent. Many cleared BP monitors are made abroad, so this is common - but each foreign establishment registers for its role and needs a valid U.S. Agent before it can complete registration and import.
13. What labeling does a blood pressure monitor need?
Device labeling under 21 CFR Part 801: a clear intended use and indications (including the population and measurement conditions the device is validated for), instructions for use, required warnings/cautions, and UDI where applicable (21 CFR Part 830). Accuracy claims must match the validation, and the labeling must not overstate what the device does. For a consumer monitor, clear, truthful instructions and accuracy information are essential.
14. What are the most common BP monitor compliance mistakes?
Marketing a monitor without the required 510(k); offering a cuffless/wearable BP feature as "wellness" without clearance; relying on a disclaimer instead of getting the device authorized; inadequate ISO 81060-2 accuracy validation; overstated accuracy claims; calling it "FDA approved"; missing UDI; and (for foreign makers) missing the U.S. Agent. Most trace back to underestimating that BP measurement is a regulated device function.
15. Is a pulse/heart-rate feature treated like blood pressure?
Not identically - different measurements have different risk and regulatory treatment. A basic heart-rate feature on a consumer wearable is often handled as a low-risk general-wellness function, whereas blood-pressure measurement is treated as a regulated device function (as the 2025 wearable action showed). Each physiologic measurement should be assessed on its own intended use and claims; don't assume a BP feature follows the same light-touch approach some heart-rate features do.
16. How does FDA Registration Assistance help BP monitor companies?
We confirm the classification (Class II, 21 CFR 870.1130, product code DXN), map the 510(k) and standards strategy (ISO 81060-2 accuracy validation, IEC 60601/EN 80601-2-30 safety, ISO 10993 biocompatibility), and handle FDA establishment registration and device listing, labeling/UDI, and U.S. Agent service for foreign makers - including for cuffless/wearable BP products where the device line is easy to cross. We're an independent compliance firm serving 1,000+ companies across 135+ countries with 15+ years of experience. Contact info@fdaregistrationassistance.com or +1 (928) 275-8333.